Faculty of Science
Faculty of Science
UNIverse - Public Research Portal

Molecular Pharmacy (Ricklin)

Projects & Collaborations

21 found
Show per page
Project cover

Discovery and Development of Drug-Like Macrocyclic Peptides for Therapeutic Complement Modulation

Research Project  | 1 Project Members

The complement system plays a crucial role in innate immunity as a first line of defense against microbial intruders. However, its dysregulation can cause or exacerbate numerous inflammatory disorders when its natural microbial elimination mechanisms “spill over” and cause damage to human cells. Therefore, therapeutic modulation of complement responses is a rapidly emerging clinical strategy, with 11 of the 12 approved drugs introduced since 2018, which covers a broad spectrum of indications - from ocular to renal and hemolytic diseases. However, various medical needs remain unaddressed. The cascading nature of the complement system, featuring an array of classically “undruggable” targets within its >30 components, favors biologics as therapeutic modalities – with restricted distribution and application profiles. This is particularly evident in neuroinflammatory diseases that, despite demonstrated associations with complement overactivation, have largely remained inaccessible for complement-targeted biologics. Moreover, many important targets within the complement cascade have proven challenging to modulate with common modalities.

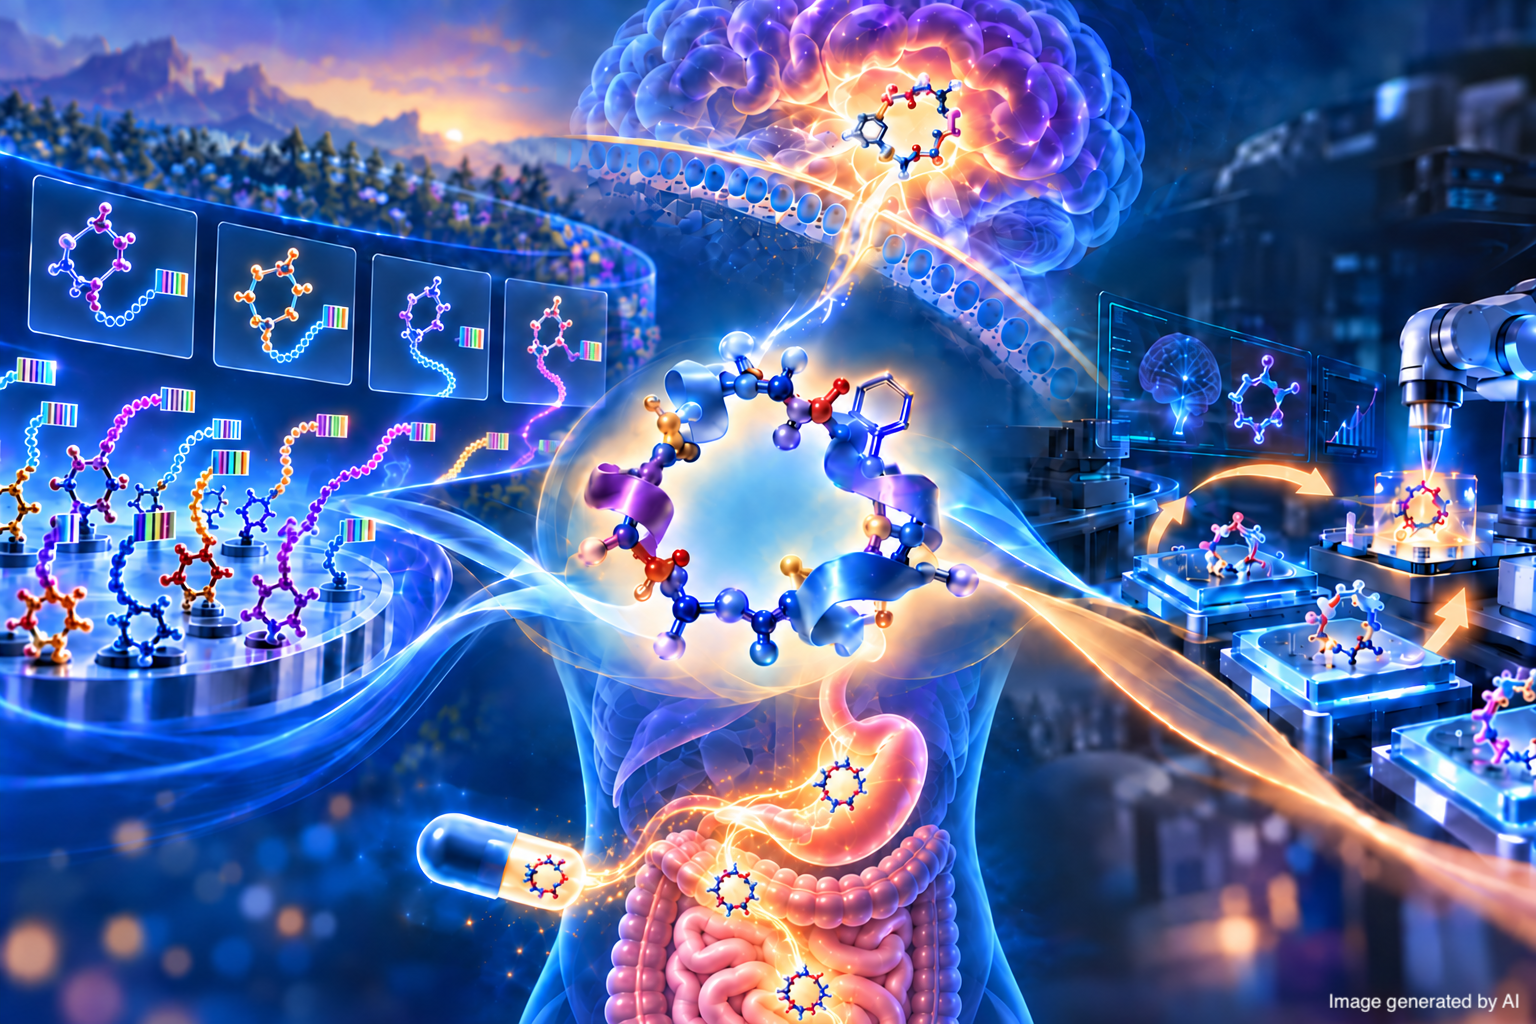
Drug-like macrocyclic peptides offer potential to address this gap – with their small, rigid-ring topology, they can engage challenging targets such as protein-protein interactions, while retaining the potential for drug-like characteristics beyond Lipinski’s “rule of 5” (bRo5), including oral bioavailability and brain penetration. Complement-targeted macrocycles may thus not only afford attractive alternatives to existing therapies by improving administration options, but may also enable complement modulation in the brain. Based on strong preliminary data, and initially focusing on established-yet-relevant targets, this proposal aims at facilitating the discovery and development of drug-like macrocyclic peptides as modulators of complement and other host defense pathways.

Upon detection of danger signals, the initiation of complement responses critically relies on the activation of serine proteases, thus serving as a bottleneck of downstream processes. While considered druggable targets, their high structural homology with other circulating serine proteases presents a challenge for small molecule development. To date, only one initiation inhibitor, the monoclonal antibody sutimlimab that inhibits the C1s protease of the classical pathway, has been approved. We previously developed small macrocyclic peptide inhibitors of C1s which serve as promising alternatives. In aim 1, we will implement high-throughput SAR campaigns to optimize these into drug-like macrocycles, with enhanced potency and desirable bRo5 properties. 

In contrast, most of the approved therapeutics are targeting central protein-protein interactions within the complement cascade, which could potentially be addressed by drug-like macrocycles. However, de novo discovery of drug-like macrocyclic peptides that can target such undefined binding pockets is challenging; fully synthetic methods that enable screening within the bRo5 space suffer from low diversity, while genetically-encoded methods deliver high diversity at the expense of the chemical flexibility needed to explore the drug-like space. In aim 2, we develop a new peptide nucleic acid (PNA)-encoded macrocyclic peptide (PeMP) screening platform. The use of PNA as the genetic barcode enables full orthogonality with modern peptide chemistry, facilitating broad chemical exploration of the bRo5 drug-like space, whilst maintaining high library diversities associated with genetic barcoding. In aim 3, we aim to apply the PeMP screening platform for the discovery of new, drug-like macrocyclic inhibitors of complement activation, although the platform is anticipated to be broadly applicable to an array of drug targets and may significantly advance de novo discovery campaigns. 

Collectively, the proposal not only aims to deliver new drug-like macrocyclic complement modulators but also aims to develop a new screening platform for discovery of drug-like macrocycles directly in hit identification – benefiting the wider research and drug discovery communities.

Project cover

Design, synthesis, and evaluation of collectin-11 inhibitors for biomedical research and therapeutic development

Research Project  | 2 Project Members

The complement system is an important innate immune pathway that plays critical roles as a first line of defense against invading organisms, damaged tissues, and the maintenance of healthy cell populations. However, its uncontrolled activation or insufficient regulation contributes to diverse clinical conditions affecting millions of patients worldwide, such as ischemia-reperfusion injury (IRI; during myocardial infarction, stroke, or organ transplantation), age-related macular degeneration, and several neurodegenerative disorders. Therefore, complement has steadily moved into the focus of drug development efforts. Whereas most therapeutic strategies focus on blocking individual effector functions, preventing the initial activation of complement could offer distinct advantages.


Collectin-11 (CL-11), a soluble C-type lectin that recognizes mannose (Man)- and fucose (Fuc)-containing glycans at the surface of pathogens or damaged cells is one such target of interest. CL-11 was recently implicated in the pathology of both acute and chronic kidney injury. In IRI-mediated acute kidney injury associated with transplantation, the expression of CL-11 is significantly elevated and co-localizes with Fuc-rich cell-surface glycans to initiate complement-dependent destruction of the kidney and loss of organ function. In chronic kidney injury, CL-11 binds to Man-based structures and acts as a leukocyte chemoattractant and enhancer of fibroblast proliferation, resulting in renal fibrosis. CL-11 also activates complement in retinal stem cells, restricting the use of stem cell transplantation for patients with age-related macular degeneration, the leading cause of blindness in the developed world. In both cell and animal models, these harmful immune responses were suppressed: (i) in CL-11 knockout mice; (ii) through enzymatic cleavage of cell-surface Man/Fuc; or (iii) by treatment with high concentrations of soluble Man/Fuc. While high-dose Fuc therapy largely prevented complement-mediated tissue damage in mice, this treatment option has real-world limitations concerning specificity, dose requirements, and pharmacokinetic (PK) profile. Therefore, the availability of CL-11-specific inhibitors with improved efficacy and drug-like properties is of therapeutic interest and would provide an important tool for delineating CL-11’s (patho)physiological role in disease.


This joint proposal between lead investigators Dr. Rachel Hevey and Prof. Martin Smieško combines their strong and synergistic expertises in computational modeling, glycomimetic drug design, chemical synthesis, and functional evaluation, to develop a novel class of specific CL-11 inhibitors. Based on the emerging role of CL-11 in renal disease and IRI, we envision that the development of carbohydrate-based, selective inhibitors for CL-11 could eventually provide a promising therapy for preventing or reducing tissue damage. To our knowledge, there are no active development programs for glycomimetic inhibitors of CL-11, and therefore the development of molecules with high affinity, selectivity, and suitable PK properties constitutes an innovative project with a potential for real-world impact.

Project cover

Mucin O-glycans as regulators of pathogen virulence: glycan synthesis, mechanistic investigation, and therapeutic potential

Research Project  | 3 Project Members

Current estimates suggest that nearly 1.3 million lives are lost annually to antibiotic-resistant infections, with this number rising to 10 million by 2050. Antibiotic resistance results in failed treatment, prolonged hospital stays, and increases the financial burden placed on medical systems and society. Therefore, novel approaches to antibiotic use are urgently needed.


Recently, a novel class of anti-virulence compounds based on mucosal glycans were discovered, which are active against a range of pathogenic cross-kingdom species, including fungal, Gram-positive, and Gram-negative bacterial pathogens. These mucin O-glycans do not directly affect pathogen survival but instead repress virulence, thereby increasing susceptibility to host immune pathways and indirectly reducing infection load. By exerting their activity through virulence attenuation instead of broad-spectrum elimination, these compounds can promote the reestablishment of a healthy microbiome and are expected to be at a reduced risk of developing drug resistance. Although this exciting new class of compounds has demonstrated broad activity, the mechanism(s) by which these glycans act has not yet been elucidated. By identifying their discrete molecular target(s), specific binding interactions can be analyzed and more drug-like glycomimetic therapies can be developed.


Mucins contain hundreds of different O-glycans which are not commercially available, cannot be purified as single structures from native sources, and are not readily amenable to solid-phase synthesis. Therefore, in ongoing work we have been developing a platform to generate individual mucin glycan structures and evaluate their ability to regulate pathogen virulence. In the further development and application of this platform, specific aims of the current proposal are focused on: (i) the recombinant expression of predicted virulence-regulating proteins; (ii) the investigation of mucin glycan binding to virulence-regulating proteins through parallel techniques; and (iii) further expansion of the mucin glycan library and the development of next-generation glycomimetic molecules.


The proposed project will improve our understanding of mucus–pathogen relationships, provide insights into virulence-attenuating mechanism(s), and provide the next steps to development of an innovative therapeutic.

Project cover

Leech-Derived Bivalent Peptide Inhibitors of Complement Initiating Proteases

Research Project  | 2 Project Members

The complement system plays an essential role in host immunity, assisting in the recognition and clearance of pathogens and damaged cells. However, dysregulation of the complement system can lead to tissue damage, contributing to a variety of inflammatory diseases. Over recent years, the therapeutic modulation of complement activity has emerged as a promising clinical intervention, creating an urgent need for new modes of complement regulation and therapeutic candidates. This proposal takes inspiration from nature to investigate the development of a new class of complement regulators.


Evolved to avoid innate immune responses, the giant Amazon leech secretes gigastasin, a protein capable of blocking the proteases responsible for initiating the complement cascade. However, the application of gigastasin as a therapeutic candidate is hampered by the complexity in its recombinant production, and the potential for triggering undesired immunogenicity. An alternative approach is to utilize the structural and mechanistic information of protease inhibition by gigastasin, to derive new molecules capable of mimicking the key target interactions. Peptides are attractive candidates as they can maintain the critical protein-target interactions, but can be accessed by scalable and modular chemical synthesis, while also minimizing immunogenicity.  


In this proposal, peptides will be derived from distinct portions of gigastasin that bind to the active center and exosite of the protease, and covalently linked together to create bivalent peptide inhibitors. The resulting peptides will be assessed for target affinity, protease inhibition, and subsequently, the impairment of complement activation in vitro. Furthermore, gigastasin, from which the peptides will be derived, exhibits multi-target protease inhibition, including those involved in both complement and coagulation pathways. Taking advantage of the highly modular chemical synthesis, structure guided optimization of the peptides towards each of the target proteases will be conducted by employing both canonical and unnatural amino acids, going beyond the current state-of-the-art in host defense protease regulation. Finally, the translational potential of the resulting peptide candidates will be evaluated, testing for complement modulation in physiologically relevant models, such as hemolytic assays, and complement inhibition in human and non-human serum samples.  


The development of leech-derived bivalent peptide inhibitors in this proposal will provide critical insights into complement protease regulation, not only providing tools for selective pathway modulation in research, but also improving our understanding of target selectivity tuning employed by parasites. Motivated by the growing success of peptides in clinical complement modulation (pegcetacoplan, Empaveli/Syfovre, Apellis; Zilucoplan, Zilbrysq, UCB, inc.), the resulting peptide candidates may also provide a platform for future therapeutic evaluation and development.